Computational modelling of kinetics and thermodynamics of eco-friendly dimerization of cyclopentadiene
DOI:
https://doi.org/10.33003/chemclass-2026-1002/13
Keywords:
Computational Chemistry; Dimerization; Cyclopentadiene; Density Functional Theory; Thermodynamics; KineticsAbstract
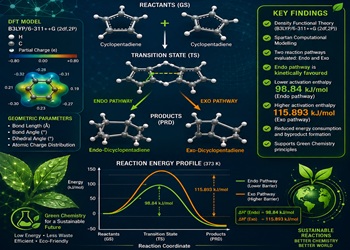
This study explores the application of computational modeling in optimizing the kinetics and thermodynamics of the dimerization of cyclopentadiene, a fundamental organic reaction with relevance in polymer and pharmaceutical industries. Using Density Functional Theory (DFT) at B3LYP level with 6-311++G (2df, 2P) basis set in Spartan software. Geometric parameters such as bond length, bond angle and dihedral angle, atomic charge distribution for the reactant (GS), transition state (TS) and the product (PRD) were obtained and activation parameters were calculated at 373K. The study evaluates reaction pathways which show endo- and exo-pathways that minimize energy consumption and byproduct formation, aligning with the principles of green chemistry. A lower activation enthalpy (98.84KJ/mol) means the endo pathway has a lower energy barrier and kinetically favored compared with the activation enthalpy for exo-pathway (115.893 kJ/mol) which indicates that the exo-pathway requires more energy to proceed, making it slower than the endo pathway under the same conditions. In conclusion, this computational DFT study demonstrates that the endo pathway for cyclopentadiene dimerization is kinetically preferred due to its lower activation enthalpy (98.84" kJ/mol" ) compared to the exo pathway (115.893" kJ/mol" ), offering valuable insights for optimizing reaction conditions at 373 K. These results highlight efficient, low-energy routes that reduce byproduct formation.Downloads
Download data is not yet available.
Downloads
Published
2026-06-30
Issue
Section
Original Research Articles
License

This work is licensed under a Creative Commons Attribution 4.0 International License.
All the work in this journal are licensed under a Creative Commons Attribution 4.0 International License.
How to Cite
Computational modelling of kinetics and thermodynamics of eco-friendly dimerization of cyclopentadiene. (2026). ChemClass Journal, 10(2), 89-99. https://doi.org/10.33003/chemclass-2026-1002/13



 © 2026 ChemClass Journal